If you plan to conduct clinical trials of medicinal products in the European Union/European Economic Area, the Clinical Trial Information System (CTIS) is an indispensable core platform. As an online system supporting the EU Clinical Trial Regulation (CTR), it undertakes all functions for clinical trial application, approval and full‑life‑cycle supervision. Starting from today, Beijing Bailian Da Pharmaceutical Technology Co., Ltd. presents a series of regulatory interpretations on CTIS.
When is CTIS mandatory?
Under the Clinical Trial Regulation (CTR), sponsors must submit applications via CTIS to obtain trial authorisation for any clinical trial involving investigational medicinal products to be carried out in the EU or European Economic Area.
Two special scenarios:
Non‑interventional (observational) studies: Mandatory submission through CTIS is not required; voluntary registration in the HMA‑EMA Real‑World Study Inventory is available.
Clinical trials for medical devices: These are governed by local medical device regulations of target countries and are not subject to CTIS.
Access Points for CTIS
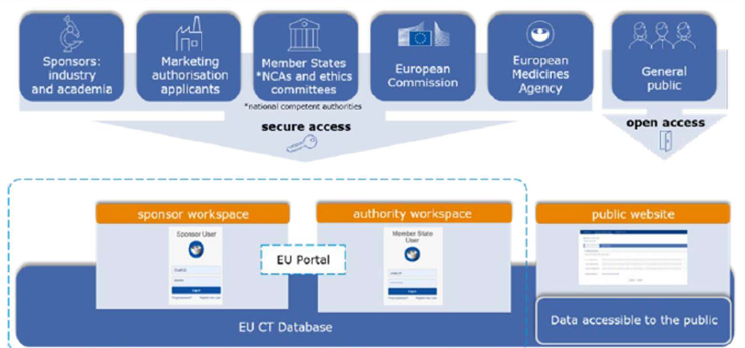
CTIS consists of two restricted secure workspaces plus a public portal website:
Sponsor workspace: Used by pharmaceutical enterprises, academic institutions and marketing‑authorisation applicants.
Authority workspace: Used by national competent authorities (NCAs), ethics committees, the European Commission and the European Medicines Agency (EMA).
Public website: Open to all users without registration for accessing public clinical‑trial information.
Preparations Prior to First‑time CTIS Use
A valid EMA account is required.
The sponsor shall have completed registration in the Organisation Management Service (OMS).
The sponsor must complete EudraVigilance (EV) registration for subsequent XEVMPD entries.
The sponsor shall ensure that information relating to investigational medicinal products for the trial has been entered into the extended EudraVigilance Medicinal Product Dictionary (XEVMPD).
After completing organisational registration, select one of the two user‑management modes below:
Organisation‑centric mode: Suitable for sponsors conducting clinical trials on a regular basis. Super administrators are designated to centrally manage roles and permissions for all personnel within the organisation.
Trial‑centric mode: Suitable for sponsors conducting clinical trials on an occasional basis. No organisational administrator is required; permissions are assigned at the individual‑trial level by the user who creates the application.
Key Takeaways
CTIS is more than merely a submission gateway; it serves as the regulatory foundation covering the full life cycle of EU clinical trials for medicinal products.
Beijing BrilliantMed Pharma Tech Co., Ltd. provides Chinese pharmaceutical companies with one‑stop services including EU clinical‑trial application support, SPOR organisational registration assistance, enterprise‑level CTIS system implementation support, global eCTD publishing and submission, and registration strategy planning. We help enterprises avoid practical pitfalls related to CTIS permissions, transparency rules and trial migration, supporting compliant market entry into Europe for Chinese pharmaceutical firms.